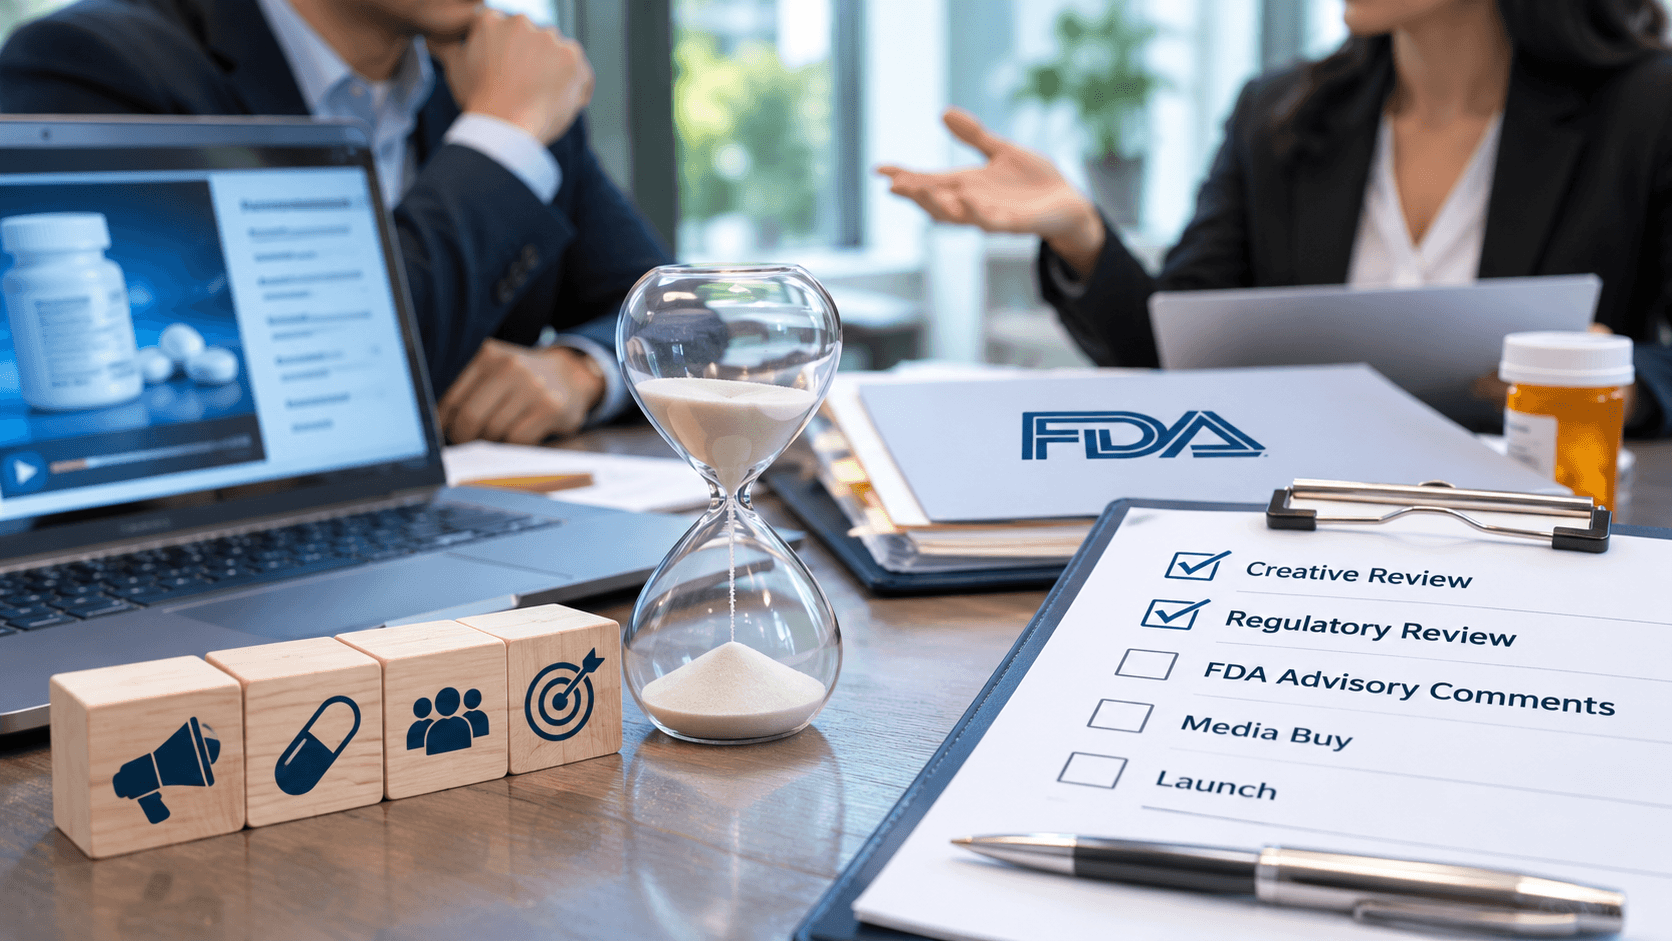
Thirty FDA enforcement letters in one year raise an uncomfortable question for drug marketers: how many campaigns could have been fixed before the first impression? During fiscal 2026, the agency’s Office of Prescription Drug Promotion (OPDP) challenged ads across multiple channels. Drugmakers can request FDA promotional advisory comments on draft ads, but launch pressures often make that choice harder than it sounds. That sounds like a useful safeguard. Yet launch calendars rarely leave room for uncertainty. For brand leaders, the question is when the benefit of agency feedback outweighs the cost of waiting.
Table of Contents
- What FDA’s 30 Letters Reveal About Ad Risk
- Why Pharma Teams Skip Prelaunch FDA Feedback
- How OPDP Review Timing Changes Launch Plans
- When Regulatory Consultation Is Worth the Delay
- Conclusion
- FAQ
What FDA’s 30 Letters Reveal About Ad Risk
At an October 28, 2026, industry conference, OPDP director Twyla Mosey reported 30 enforcement letters during fiscal 2026: 28 untitled letters and two warning letters. According to FDA Webview’s account, she said a meaningful number might have been prevented through greater use of prelaunch consultation. OPDP also received more than 100 advisory-comment requests and over 172,000 promotional materials filed on Form FDA 2253. However, those figures reflect different processes, not an approval rate. For most prescription drugs, submitting final advertising at first use is required, while requesting feedback on drafts is voluntary.
The violations involved more than awkward wording. FDA challenged minimized safety risks, unsupported efficacy claims, misleading comparisons, and inaccurate descriptions of approved uses. Meanwhile, television ads, social posts, brochures, and conference displays all drew scrutiny. A brand might satisfy an internal checklist yet still create a misleading overall impression.
Consequently, the question is whether anyone can challenge a campaign’s central message before production and media commitments make changes painful. Early intervention may prevent more than a regulatory headache. It can also protect campaign budgets, brand credibility, and relationships with healthcare professionals.
Why Pharma Teams Skip Prelaunch FDA Feedback
Early feedback from the FDA can identify potential problems while campaign materials are still easy to change. However, seeking that feedback creates a tradeoff with speed. Commercial teams may be coordinating agency shoots, medical-legal-regulatory reviews, sales training, and media inventory at once. Each has its own deadline and budget. Delaying one asset can disrupt an entire launch plan or an important competitive window.
Timing is especially costly in crowded treatment markets. A rival may reach patients first, while paid media reservations expire or production crews become unavailable. Moreover, marketing teams may believe their internal review already tested every claim. That belief can grow stronger after months of approvals.
Yet an internal committee cannot guarantee that FDA will interpret a tagline or image the same way. As Pharma Marketing Network’s analysis of fair balance explains, the combined effect of visuals, sequencing, and claims matters as much as individual disclosures.
There is also a financial calculation. Companies may compare the uncertain cost of regulatory scrutiny with the immediate expense of postponing a campaign. However, that calculation can overlook the cost of replacing creative assets, revising media plans, or responding to enforcement concerns after launch. Therefore, skipping consultation may preserve short-term momentum while leaving a larger risk unresolved.
How OPDP Review Timing Changes Launch Plans
FDA does not routinely preapprove most prescription drug ads. Under its OPDP guidance, companies generally submit final promotional materials when first published or distributed, while requesting opinions on drafts is a separate option. The distinction matters because filing Form FDA 2253 does not mean the agency reviewed an ad before launch.
For core introductory launch materials, OPDP has a 45-calendar-day response goal, but that is not a guaranteed end-to-end deadline. A five-business-day screening comes first to check whether a submission is complete and properly annotated. In addition, consultation with other FDA experts can extend the timeline, while noncore materials receive lower priority.
Therefore, a submission made shortly before a major media buy may not protect the planned airdate. Marketers should work backward from launch, identify claims requiring additional expertise, and supply clear references. Early preparation reduces the chance that review time becomes a commercial emergency.
However, simply adding 45 days to a campaign schedule may not be enough. Teams should allow time for internal revisions, possible follow-up questions, and any required creative changes. This approach helps avoid a situation in which a campaign is technically ready for distribution but commercially difficult to modify.
More importantly, regulatory planning should begin before agencies finalize scripts, storyboards, and media placements. When compliance is treated as part of campaign development rather than a final checkpoint, teams have more flexibility to respond to concerns without disrupting the entire launch.
When Regulatory Consultation Is Worth the Delay
Asking the FDA to review draft promotional materials makes the most sense when a misleading claim could lead to costly changes or enforcement action. Consider a new efficacy message, a comparison with a competitor, a complex safety claim, or a prominent consumer video. Those elements can be costly to correct after production and distribution are locked.
In contrast, a familiar claim backed by strong evidence may call for rigorous internal review rather than another external step. The goal should not be to send every promotional asset to the agency. Instead, companies need a practical way to identify campaigns where additional scrutiny provides meaningful value.
Commercial and regulatory leaders can establish a risk-based decision point before approving creative. Ask what each claim implies, whether evidence supports that impression, and what it would cost to replace the asset. Next, decide whether agency feedback belongs on the critical path or whether the concept should be simplified.
Teams also need a backup plan if comments arrive after a tentative launch date. That might involve holding a high-risk message, adjusting the media schedule, or preparing alternative creative that relies on established claims.
As Pharma Marketing Network’s report on conference booth compliance shows, regulatory scrutiny extends well beyond television. Digital campaigns, social media promotions, and conference materials can also raise concerns about misleading product communications.
Ultimately, the strongest launch schedule leaves room to fix the message before buying reach. Regulatory consultation should be viewed as a strategic investment when the potential cost of an advertising mistake exceeds the commercial value of moving faster.
Conclusion
Getting feedback from the FDA does not guarantee compliance or protect a company from future enforcement. Still, the fiscal 2026 enforcement record suggests some avoidable problems survived routine launch workflows.
For marketers, the choice should not be asking FDA about every banner or skipping the process entirely. Instead, schedule consultation early for high-risk campaigns, protect time for revisions, and keep defensible claims at the center.
A little more planning may cost less than pulling a campaign after it reaches the public. The real opportunity is to make regulatory consultation part of smarter commercial decision-making rather than treating it as another obstacle to launch.
FAQ
Does the FDA require drugmakers to submit ads for review before launch?
For most prescription drug materials, prelaunch advisory review is voluntary. However, certain products, including those approved under accelerated approval regulations, face separate presubmission requirements. FDA may also require preapproval in specific circumstances.
Does advisory feedback mean FDA approved the ad?
No. OPDP provides opinions on proposed promotional communications, not a blanket promise against future enforcement. Drugmakers remain responsible for ensuring their advertising complies with applicable regulations.
How long should a brand allow for OPDP review?
Core launch reviews have a 45-calendar-day response goal following initial screening. However, consultations with other FDA experts can extend the total review time. Companies should avoid treating the target as a guaranteed deadline.
What happens if a brand skips FDA feedback?
Most campaigns can proceed after appropriate internal review and required submissions. However, the company remains responsible for its promotional content and could face an FDA enforcement letter if the advertising is misleading or otherwise violates applicable requirements.
When should pharmaceutical marketers consider requesting FDA feedback?
Prelaunch consultation may be particularly valuable for complex efficacy claims, comparative advertising, significant safety messages, or campaigns involving substantial media investments. Early evaluation can help teams determine whether the regulatory benefit justifies the additional review time.
This content is not medical advice. For any health issues, always consult a healthcare professional. In an emergency, call 911 or your local emergency services.